

What's Happening?
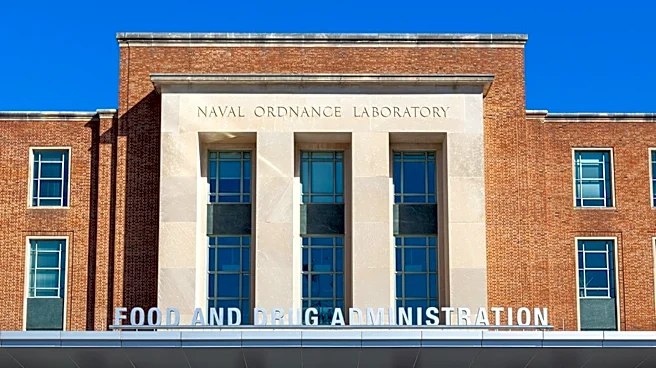
The U.S. Food and Drug Administration (FDA) has released new draft guidance that supports the use of Bayesian methods and simulation-based evidence in clinical trials. This guidance signifies a notable shift in the regulatory approach to predictive medicine.
Dr. Irina Babina, CEO of oncology R&D platform Concr, highlighted that existing regulatory frameworks were primarily designed for static, single-test diagnostics. She emphasized that if the FDA continues to approve AI medical tools using the same criteria as traditional blood tests, the most promising predictive technologies may not reach patients. The new guidance aims to modernize the evaluation process for advanced medical technologies, particularly those leveraging artificial intelligence and complex data analysis.
Why It's Important?
This FDA draft guidance is crucial for the advancement of predictive medicine and personalized healthcare in the U.S. By endorsing Bayesian methods and simulation, the FDA is acknowledging the potential of more sophisticated statistical approaches that can provide more robust and efficient clinical trial designs, especially for complex diseases like cancer. This could accelerate the development and approval of innovative treatments and diagnostic tools. For the pharmaceutical and biotechnology industries, this guidance offers a clearer pathway for integrating AI and advanced analytics into their R&D processes. However, it also underscores the challenge of adapting regulatory frameworks to keep pace with rapid technological advancements, ensuring that groundbreaking technologies are not hindered by outdated evaluation criteria. The shift could ultimately benefit patients by bringing more effective and tailored medical solutions to market faster.
What's Next?
The draft guidance will likely lead to further discussions and refinements as stakeholders, including pharmaceutical companies, AI developers, and medical professionals, provide feedback to the FDA. The oncology R&D platform Concr, led by Dr. Irina Babina, is actively engaged in these discussions, advocating for regulatory approaches that are better suited for predictive technologies and digital twins. The FDA's move suggests a future where clinical trials could become more adaptive and data-driven, potentially reducing the time and cost associated with drug development. Companies developing AI-powered diagnostics and therapies will need to align their development and validation strategies with these evolving regulatory expectations to ensure successful market entry. The long-term impact could be a more dynamic and responsive regulatory environment that fosters innovation in medical technology.
Beyond the Headlines
The broader implication of this FDA guidance extends to the ethical and practical considerations of integrating AI into healthcare. The concept of 'digital twins' – virtual models of patients used for simulation – raises questions about data privacy, algorithmic bias, and the interpretability of AI-driven predictions. Ensuring that these advanced technologies are not only effective but also equitable and transparent will be paramount. Furthermore, the shift towards simulation-based evidence could redefine the standards of 'proof' in medicine, moving beyond traditional randomized controlled trials in certain contexts. This could lead to a more nuanced understanding of disease progression and treatment response, but also necessitates robust validation methods to maintain patient safety and trust. The guidance also highlights the ongoing tension between rapid technological innovation and the need for rigorous regulatory oversight.