


What's Happening?
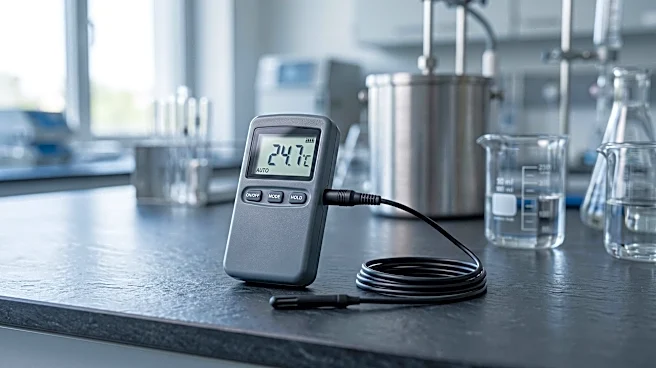
Physitemp Instruments, LLC has initiated a Class 2 device recall for its Physitemp MT-D temperature monitors and associated probes. The recall, initiated on September 28, 2026, and posted by the FDA on November 10, 2026, affects all lot codes distributed
between January 19, 2024, and January 19, 2026. The manufacturer's reason for the recall is that these temperature monitors and probes have not been reviewed for safety and effectiveness for their intended use. The FDA determined the cause to be 'No Marketing Application.' Physitemp Instruments has issued an 'Urgent: Medical Device Correction' notification, instructing consignees to label these devices as 'ARUO' (Animal Research and General Non-Human Laboratory Purposes Only) and to discontinue any human clinical use immediately. The recall impacts 36 units distributed nationwide across 20 U.S. states and internationally.
Why It's Important?
This Class 2 recall of Physitemp MT-D temperature monitors is significant because it highlights a critical lapse in regulatory compliance and patient safety. The devices were distributed without proper review for safety and effectiveness for their intended use, which could potentially lead to inaccurate temperature readings or other malfunctions if used in human clinical settings. While the recall specifies that the devices are now designated for animal research and non-human laboratory purposes only, their prior distribution without clear usage limitations could have put patients at risk if they were inadvertently used in human care. The FDA's classification of this as a Class 2 recall indicates that the use of these products may cause temporary or medically reversible adverse health consequences, or that the probability of serious adverse health consequences is remote. This incident underscores the importance of rigorous pre-market evaluation and clear labeling for medical devices to ensure patient safety and proper application.
What's Next?
Physitemp Instruments, LLC has instructed affected consignees to take several immediate actions. These include reviewing internal equipment records, notifying relevant laboratory personnel, applying 'ARUO' labels to the devices, distributing updated 'Instructions for Use' inserts, and discarding legacy documentation. Crucially, any use of these devices for human clinical purposes must be discontinued immediately. Consignees are also required to sign and return an acknowledgment form within 30 days to verify receipt of this documentation update. The FDA will continue to monitor the recall process until it is terminated, ensuring that all necessary corrective actions are taken and that the devices are appropriately restricted to their designated uses. This event may lead to increased scrutiny of device marketing and distribution practices to prevent similar issues in the future.
Beyond the Headlines
The Physitemp MTD recall points to a broader issue within the medical device industry regarding the clarity of intended use and regulatory pathways. Devices initially developed for research or non-human applications can sometimes find their way into clinical settings without undergoing the necessary human-use safety and efficacy reviews. This 'mission creep' can create significant risks, as the performance and safety profiles for animal research may not translate directly to human patient care. The FDA's 'No Marketing Application' determination suggests a failure to secure the appropriate clearances for human use, highlighting a potential gap in how some manufacturers categorize and market their products. This incident could prompt a re-evaluation of how medical devices are classified and how their intended use is communicated and enforced throughout their lifecycle, particularly for products that could be ambiguously applied across research and clinical domains.