

What's Happening?
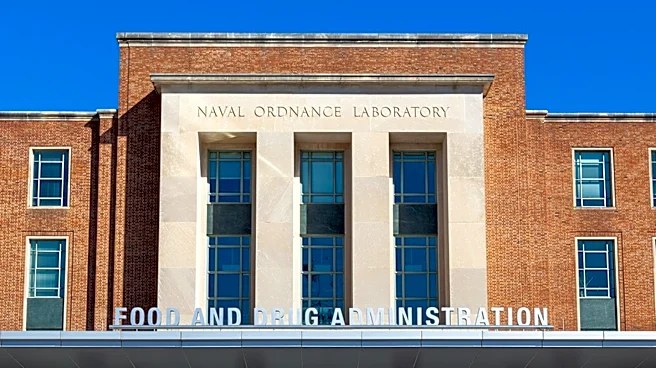
The U.S. Food and Drug Administration (FDA) has issued new guidance documents to address the evolving regulatory landscape for cell and gene therapies (CGTs). In January 2024, the FDA released guidance titled 'Human Gene Therapy Products Incorporating
Human Genome Editing,' establishing a framework for evaluating risks associated with these products. More recently, the FDA issued draft guidance focusing on next-generation sequencing (NGS) approaches for safety assessment. These documents emphasize a risk-based approach to characterizing intended editing outcomes, unintended editing, and broader genomic integrity consequences. The FDA's focus is on ensuring comprehensive assessment of factors such as on-target editing, off-target effects, structural rearrangements, and vector-related integration, especially as the field moves beyond CRISPR-Cas9 applications to more diverse editing strategies.
Why It's Important?
This evolving regulatory framework is crucial for ensuring the safety and efficacy of rapidly advancing gene therapies in the U.S. The FDA's guidance provides clarity for developers on how to design genomic safety programs that are scientifically justified and appropriate for their specific products. By focusing on a risk-based assessment, the FDA aims to mitigate potential dangers such as off-target edits, unintended nucleotide substitutions, and large structural variations that modern genome editing systems can introduce. This structured approach helps build a robust body of evidence for investigational new drug (IND) and biologics license application (BLA) packages, ultimately protecting patients while fostering innovation in a field that offers significant promise for treating genetic diseases with limited or no effective options.
What's Next?
CGT developers will need to integrate these new FDA guidelines into their early therapeutic development processes. This includes designing genomic safety strategies that are staged and risk-based, selecting complementary methods to address plausible risks, and clearly explaining assay limitations and bioinformatics parameters. The FDA's emphasis on next-generation sequencing (NGS) means developers will need to adopt advanced sequencing strategies and robust data interpretation. The regulatory landscape will continue to evolve, requiring ongoing adaptation from developers. The FDA also issued draft guidance in February 2026 on a 'plausible mechanism framework' for individualized therapies, indicating a continued focus on tailored approaches for ultra-rare diseases, which will further shape future development and regulatory submissions.
Beyond the Headlines
The FDA's proactive stance on regulating gene therapies reflects a broader societal challenge: how to balance rapid scientific advancement with patient safety and ethical considerations. The guidance documents highlight the complexity of genomic safety, moving beyond simple off-target analysis to a comprehensive evaluation of genomic integrity. This shift acknowledges that gene editing is not without risks, and a thorough understanding of these risks is paramount. The development of individualized therapies, while promising for rare diseases, also raises questions about the feasibility of traditional clinical trials and the need for flexible regulatory frameworks. This ongoing dialogue between scientific innovation and regulatory oversight will shape the future of medicine, influencing not only how diseases are treated but also how genetic information is understood and manipulated responsibly.